Spinal Muscular Atrophy (SMA) is a rare genetic disease marked by progressive muscle wasting and motor neuron loss. Variants in the SMN1 gene, which encodes the SMN protein essential for motor neuron survival, drive the disease, while the SMN2 gene acts as a modifier, giving SMA its wide range of severity and manifestations.
JAX offers a well-validated portfolio of SMA-relevant in vivo mouse models, paired with extensive preclinical service expertise to execute your SMA study goals. Partner with JAX and get deep knowledge of SMA models and Ph.D.-level study directors on every project, delivering thoughtful, comprehensive project design and execution for your research.
In addition to SMA mouse models, The Jackson Laboratory also offers human induced pluripotent stem cell (hiPSC) lines carrying neuromuscular disease-relevant variants, providing complementary tools for translational research.
See all SMA Mouse Models at JAX
| Common Strain Name | Phenotype | Survival |
|---|---|---|
Delta 7 Model* |
| Median survival of 13 days with longest observed survival of 22 days (for triple homozygous mice) |
Li Model* |
| Median survival of 13 days with longest observed survival of 17 days |
SMN1c/c* |
| Lifespan of 60+ weeks |
*Requires a license.
Partner with JAX Preclinical Services to offload the full complexity of your SMA efficacy studies. JAX brings scientific expertise and expert colony management, with study endpoints including but not limited to the following:
Speak to a JAX Study Consultant
Study variables differ depending on your model of choice and research needs. Below is a sample study:
| Timeline | 12+ days but can vary depending on study model |
|---|---|
| Groups | Typical n per group: n=10 |
| Dosing |
|
| In vivo Readouts |
|
| Post-mortem |
|
Delta 7 SMA Model (005025)
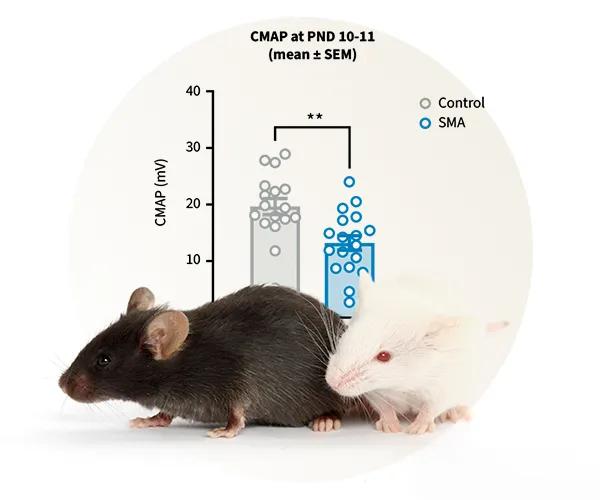
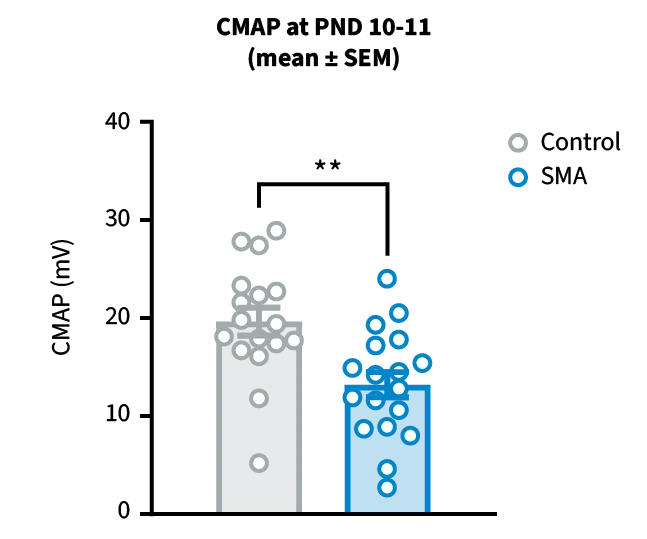
Compound muscle potential (CMAP) from the tibialis anterior muscle is decreased at P10 in the Delta 7 mice.
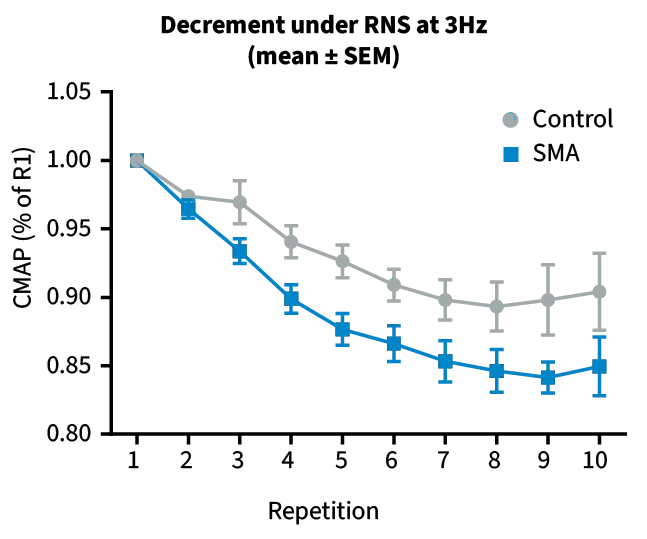
Upon Repetitive Nerve Stimulation (RNS), CMAP response declines faster in the Delta 7 mice than in the controls at P10.
Li Model (005058)
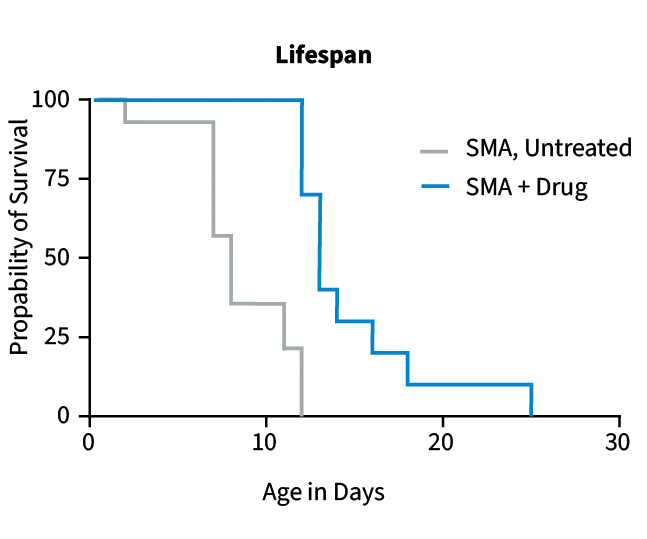
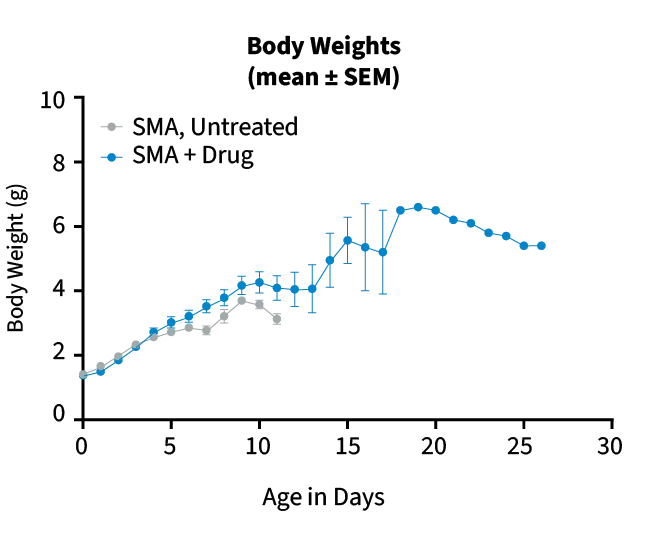
Median survival of 8 days in untreated SMA mice is extended (p<0.001) to 13 days after one neonatal injection of the test article.
Body weight starts to decline at 10 days in the SMA mice but weight gain is extended to 20 days in the treated mice.
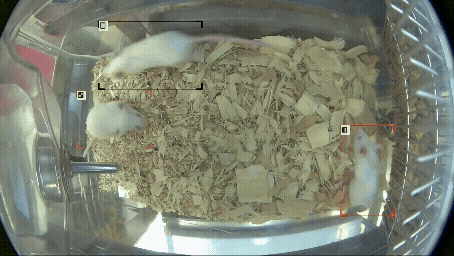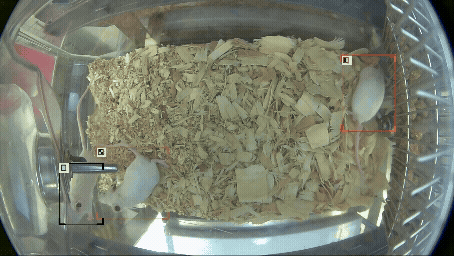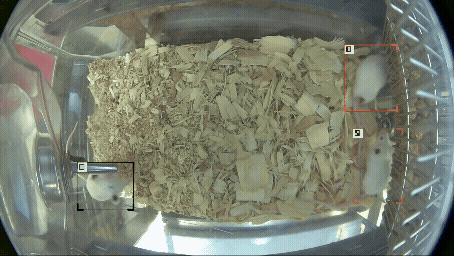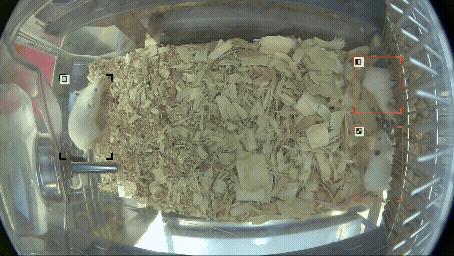
With 24/7 monitoring and AI-enhanced digital measures, spot phenotypic indicators in your research models earlier than episodic observation ever could. JAX Envision reveals day-to-day changes in an individual mouse's activity and total distance traveled, helping you catch disease indicators sooner than ever before.
This manual summarizes The Jackson Laboratory's resources for spinal muscular atrophy (SMA) research, including all available models and their genetic backgrounds, to help you choose the best one for your studies. Because each mouse model reflects a different part of the disease spectrum, in both phenotype and molecular defect, understanding these differences is key to interpreting results.